MolDraw Community

M
advikmahesh04 10 karma
Student ·

Question:Done

I've been trying to use these two smile codes but it isn't working. Are there any solutions? If there is something I am doing wrong when I click import smiles, let me know. [Na+].OC([O-])=O.Cl>>OC(O)=O.[Na+].[Cl-] [Na+].[Na+].[O-]C([O-])=O.Cl>>[Na+].OC([O-])=O.[Na+].[Cl-]

▲
noelsebastianc 10 karma
Student ·

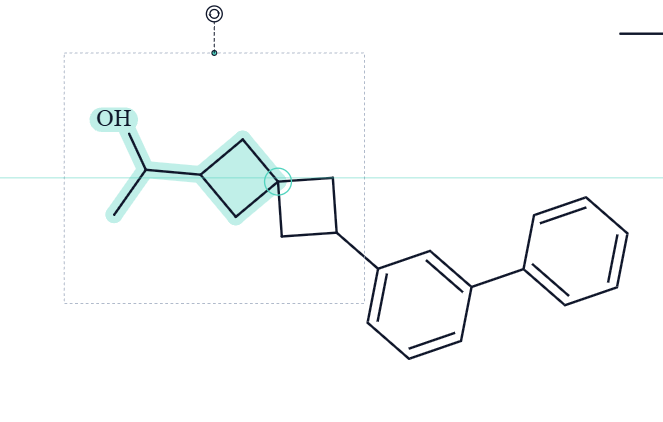
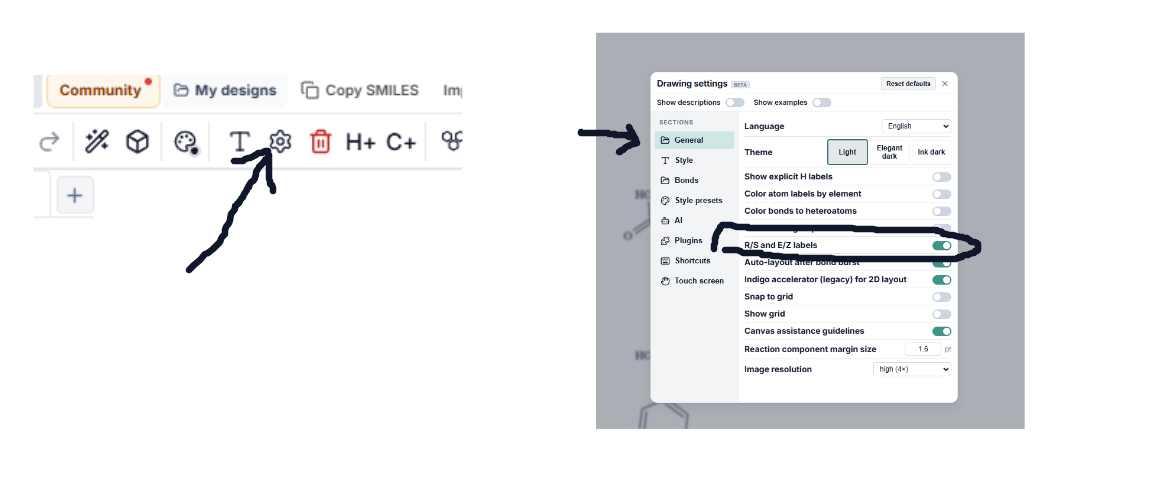
hi, how do i put and bond to representate R & S isomers?

like to add these type of bond

▲

Rafeeque

Rafeeque Admin 364 karma

Hi, To show R and S isomers in MolDraw: Draw the molecule as usual (the four groups on the chiral carbon). Select the wedge bond tool (solid wedge = coming toward you) or the dash/hash bond tool (going away from you). Click the bond from the chiral carbon to one of the groups (often H, or another substituent). MolDraw will show an R or S badge on that carbon when the stereo is clear, and the name in molecule info can include (R)/(S).

Rafeeque Admin 364 karma
Other ·

3D rotation of molecules!

You can adjust 3D rotation of molecules in Moldraw now!

▲
Rafeeque Admin 364 karma
Other ·

Arrows are improving!

You can draw nice arrows more easily now in moldraw Try and let me know what to improve.

▲
Maxim1 Janzen 35 karma
Student ·

Some bugs

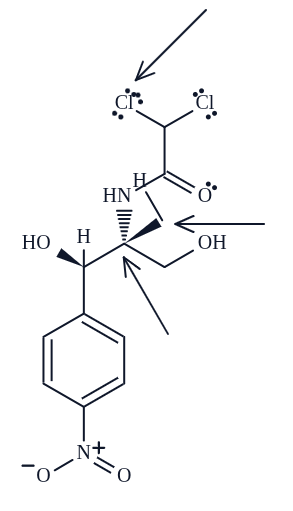
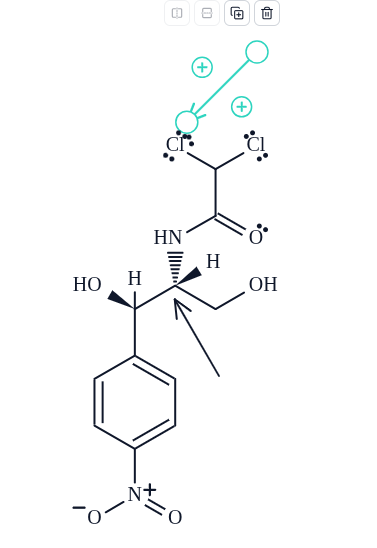
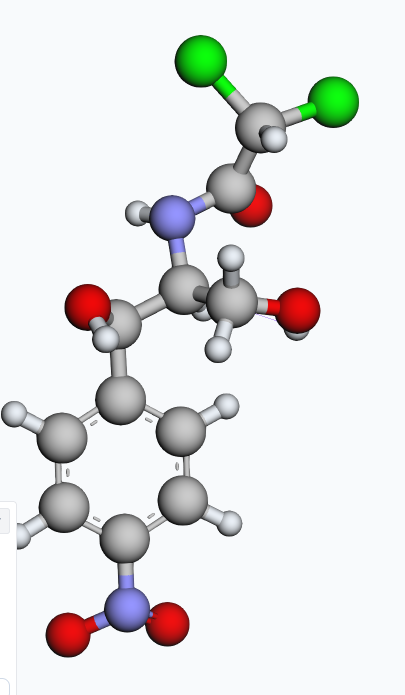
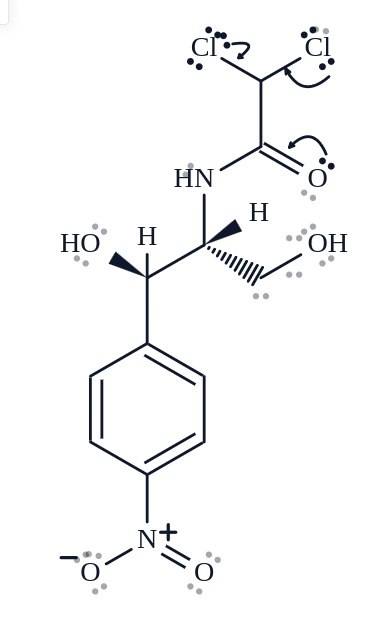
Hi, i really like your update but there is a problem with some things. First i realized now that sometimes editing/deleting posts does not work. In the pictures you can see several problems: - wrong 3d structure at the chiral center when changing the wedge direction - weird LP placement of third LPs (didnt test more than 3) - a button to add one to all of them would be nice so not each atom would have to be clicked - replacing atoms with an h fails and a methyl group is added instead - some lps are drawn wrong when choosing electron flow - in electron flow mode it is not always possible to choose the middle of a bond as endpoint - when a bond is chosen as starting point and an atom as endpoint it often points wrongly which corrects itself when some atoms are deleted (no picture, but could be done on request) Then it is also slightly annoying that enter in the forum results in the post being posted immediately as it is usually used for newlines. It would be nicer if you could only post by clicking the button or have a confirmation before hand. Thanks for your dedication and continued updates. Best Maxim

Community attachmentCommunity attachmentCommunity attachmentCommunity attachment
▲

Maxim1 Janzen

Maxim1 Janzen 35 karma

Thanks a lot for fixing all of these issues. I think it is quite nice already. I noticed that you implemented the add all lone pairs feature as "add one lone pair to all", would it perhaps be possible to have "Add all lone pairs", which adds all of the possible ones to all respective atoms at once? I think that would reduce a great number of clicks that would need to be done. It would also be nice to have the same kind of view of possible lone pairs when drawing the electron flow arrows like in the last picture in this post, it was removed since the last update but i think it was quite nice. Then a few new thing, i saw that the copy smiles (it would be nice if it was possible to import smiles also) and copy svg buttons are not working at least on my linux machine with chrome and firefox. Also the advanced export button seems to be doing nothing. And when i try to do a MMFF94 optimization i get an error about openChemLib failing to load. I hope this feedback is useful for you. Thanks for the continued updates.

Rafeeque

Rafeeque Admin 364 karma

Hi Maxim, Thanks for the feedback. Very helpful. We have solved most of the things. I will update you soon when all updates are live.

Rafeeque

Rafeeque Admin 364 karma

Hi Maxim, @maxim1janzen Thanks for the detailed notes. These are in now. Please hard-refresh the site once so you are not on an older cached page. (ctrl + f4) 1) Add all lone pairs now adds every possible lone pair on every atom in one click. Add one lone pair to all is still there if you only want a single pair on each atom. 2) While you draw an electron-flow arrow, possible lone-pair sites show again, and they go away after the arrow is placed. 3) Copy SMILES and Copy SVG should work now can you check?. Import SMILES is next to Copy SMILES. 4) Advanced export (check now?) 5) MMFF94 loads OpenChemLib again, so that optimization should run (check now?)

Luis Martin+Moldraw 23 karma
Student ·

attach atoms by draggingDone

Looks nice but, now I can't dragging atoms or groups to attach them in to an existing structure.

▲
Luis Martin+Moldraw 23 karma
Student ·

attach atoms by dragging

Looks nice but, now I can't dragging atoms or groups to attach them in to an existing structure.

▲
Rafeeque Admin 364 karma
Other ·

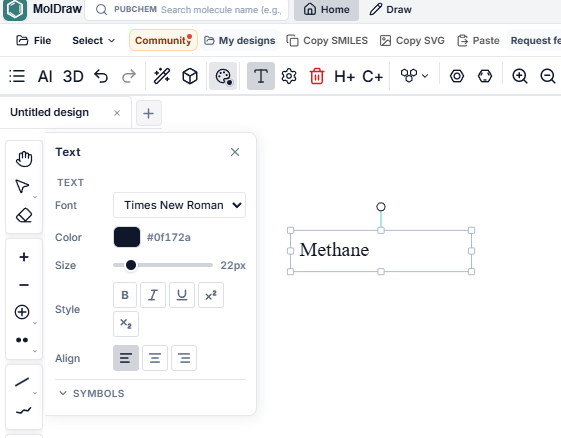
Text tool is better now!

Text tool had some issues. They are fixed. Tell me if anything to be improved further. Thanks.

Community attachment
▲
Rafeeque Admin 364 karma
Other ·

The new Moldraw version is live! 🚀

We’ve launched a major update with many fixes and new features. I know there may be some differences, bugs, or things that still need improvement. The previous editor had several limitations, so we had to make this change to build a much better experience going forward. If you face any issue, please let me know here or through the support chat. I’m personally here to fix things as quickly as possible. Give us a little time. Within the next month, I’m confident Moldraw will become a much better tool for drawing chemical structures. We’ll improve it together. 😊 Thank you for your patience and understanding!

▲

TOPHERKIRKPR

TOPHERKIRKPR 12 karma
Researcher ·

Using it again, and it's so much better than the initial update. I can make bonds again. It doesn't tell me not to make Texas carbons. The commonly used tools and notations are easier to find

Mcpsmsn 10 karma
Student ·

Chair conformations

Is there any feature here that automatically forms a chair conformation?

▲