MolDraw feature requests

M

Have an idea for MolDraw?

Request a feature here so the community can vote and track progress. Sign in to submit.

Ramon
Feature request ·

Fast switch between skeletal and lewisUnder progress

It would be great to be able to switch between skeletal formula and Lewis formula by pressing one or two buttons. Atm I can can add explicit H-Atoms and reveal C-Atoms to get to the Lewis formula. To get back to the skeletal formula I can hide the C-Atom but I cannot make the H-atoms implicit if I made them explicit before.
Rafeeque

Rafeeque Admin 368 karma

Hi, for now you can select and delete the explicit hydrogens. but we will make it easier. working on it

lawsonchioma
Feature request ·

to write names and covert to structureDone

convert written names to structure

Rafeeque

Rafeeque Admin 368 karma

Hi This is possible. right click on text > convert to structure

Feature request ·

ACSAlready implemented

ACS style

Ramon
Feature request ·

MEP Surface in 3DAlready implemented

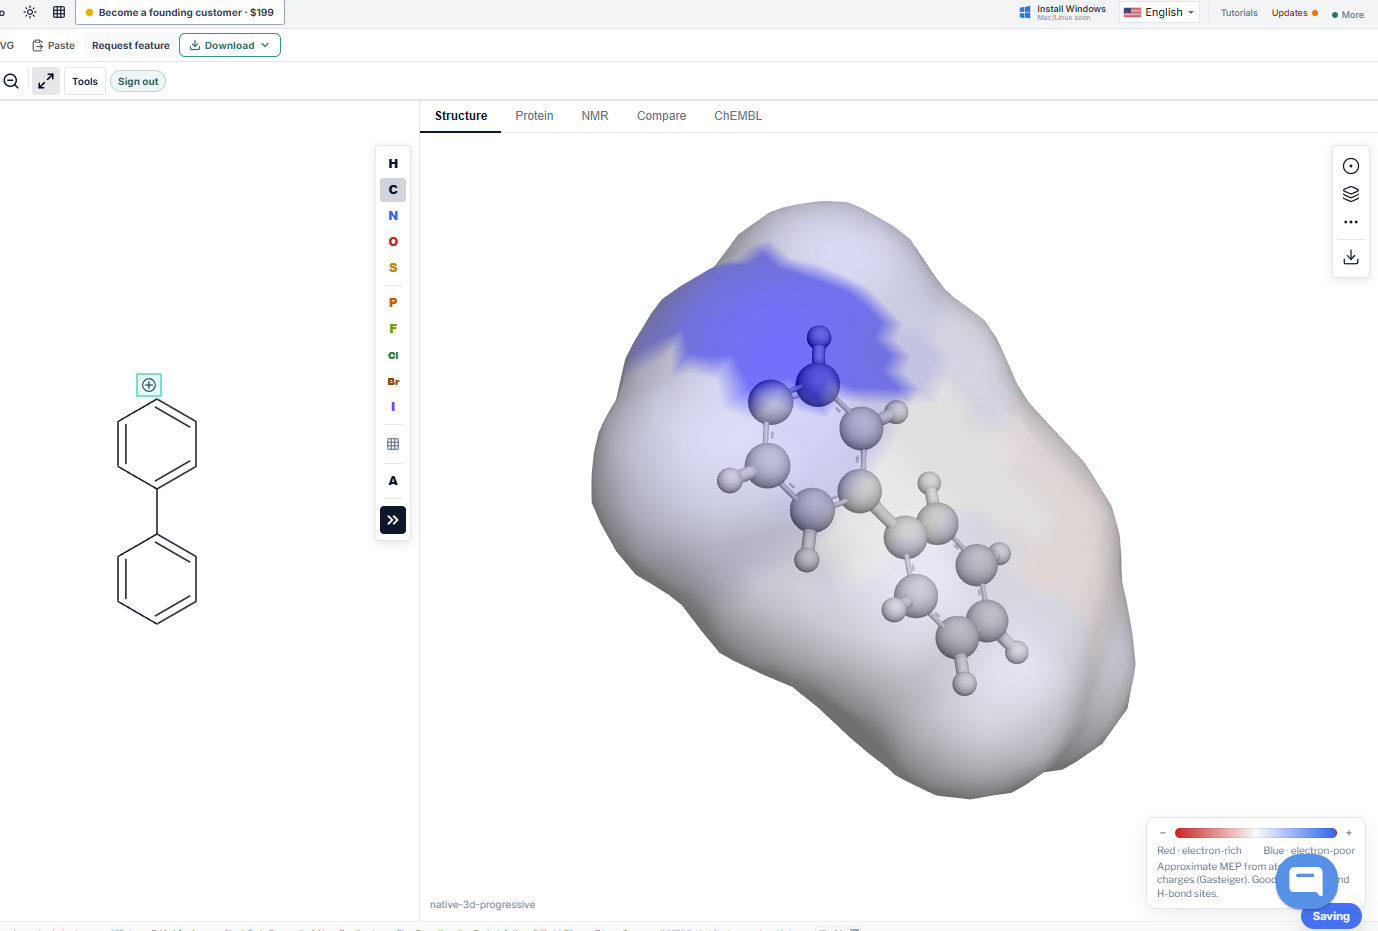
Hi there, The implementation of Molecular Electrostatic Potential (MEP) or Molecular Electric Field (MEF) for the 3D models would be great.

Rafeeque

Rafeeque Admin 368 karma

Hi @ramon A basic verison of MEP visualisation is already available.

Reply attachment

Ramon

Ramon 24 karma

I see. I didn't check the plugins. It's a bit rougher than what I used before, but it will do. Thanks.

Ramon
Feature request ·

Lone pairs as lines + movable positionsDone

MolDraw is really nice, but I miss the option to draw lone pairs as a line. Also I wasn't able to place the lone pairs in a different location like it is possible with charges. Maybe its possible to have them treated like bonds so that they take up an optimal position when using clean up and still be able to change the position to our liking. (Submitted by email after in-app feature request failed due to RLS.)

Ramon

Ramon 24 karma

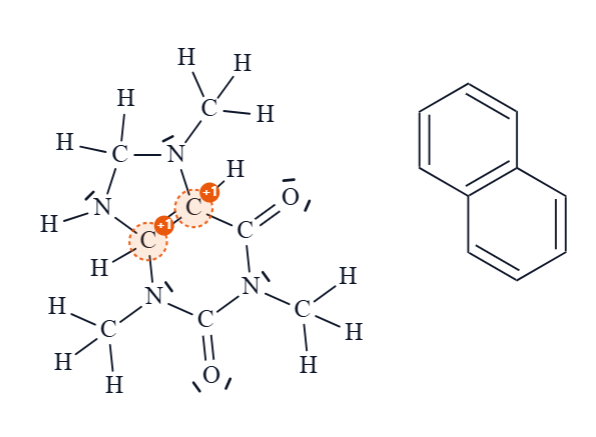
Hey Rafeeque, Great work! Checked with caffeine again. - Positions look very good. - Positions for some are locked during rotation, which is very good. I'd like it locked for all lone pairs during rotation, if possible. - Clean up will fix any bad placed lone pairs. (i.e. After rotation it is possible that a lonepair overlaps with the ring symbole of an aromat.) - On a side note: Dearomatizing caffeine doesn't work. See attached image.

Reply attachment

Rafeeque

Rafeeque Admin 368 karma

Hi Ramon, All three points are fixed and live now: 1) Lone pairs now stay fixed to their atom when you rotate. The only exception is a pair that would sit right on an OH or NH hydrogen label; it moves just off the H. 2) Clean up now moves any badly placed lone pair into the most open space, away from bonds, H labels and aromatic ring circles. 3) Dearomatize now works on caffeine and similar ring systems like purines and indole. 4) The problem in your picture came from adding explicit H while the rings were aromatic, which gave extra H atoms. That is fixed too, and Dearomatize also cleans up those extra H atoms. Please refresh the page (Ctrl+F5) and give it another try. Let me know if anything still looks off.

Ramon

Ramon 24 karma

1. Works as intended and it totally makes sense to shift them for OH and NH Labels. 2. Works as intended. 3. Works as intended. 4. Works as intended. Is there a way to get rid of all lonepairs?

View more comments
Vanessa H
Feature request ·

Font on arrows are too smallDone

Style Structure does not affect it

Feature request attachment

Rafeeque

Rafeeque Admin 368 karma

Hi Vanessa, This is fixed. Now you can adjust the font size of the text in arrows in the left settings panel whichappear when select the arrow. Also theefault font size is also increased. @vanessah

wkparaiso
Feature request ·

Search on pubchemAlready implemented

It should be able to search the structure on pubchem. Do you think it's possible?

Rafeeque

Rafeeque Admin 368 karma

Hi, You can find a search option at the top left. It is PubChem search. Is there anything else you would like to have in the pubchem search?

ahansris
Feature request ·

Coordination bondsDone

For example, for a ferrocene molecule I'd like to be able to show the dottted coordination bond lines between the iron atom and the cyclopentadienyl rings.

Webberliam23
Feature request ·

Revert back to old modelDone

Please revert back to the old format, maybe make it so that you can add lone pairs, but this new model is awful. I can't use it at all.

TOPHERKIRKPR

TOPHERKIRKPR 12 karma
Researcher ·

Reading my post back it was dramatic but I wanted to say today's update is so much better. The adding text feature is very bugged, it add colors if the color feature is on, can't use the space bar and if the text box overflows the text wraps on itself

Reply attachment

Rafeeque

Rafeeque Admin 368 karma

Thanks for the follow-up , glad today's update feels better. On the text tool: text no longer picks up the molecule Color. Space works while you type, and overflow wraps or grows the box instead of stacking on itself. Text styling is in the left panel now, the Style button opens the Text dock, and superscript/subscript apply only to the selected text. Please check and let me know ioaf nay hages are needed

Rafeeque

Rafeeque Admin 368 karma

Hi, Try this. Use Select tool, click an atom or bond to select it, then drag to move or press Delete — no need to switch to Erase each time. That solves the issue?

View more comments
NH
Feature request ·

molecule to nameDone

You draw a molecule and then the programe wil name it after IUPAC naming. and the other way around. You write the name of the molecule and then the program draws it

Rafeeque

Rafeeque Admin 368 karma

Hi, You can find a search option at top left. There you can type and search name of the molecule it will draw the structure. Select any molecule and open info box at top right (look for info option) it will open adetails box. It will have IUPAC name