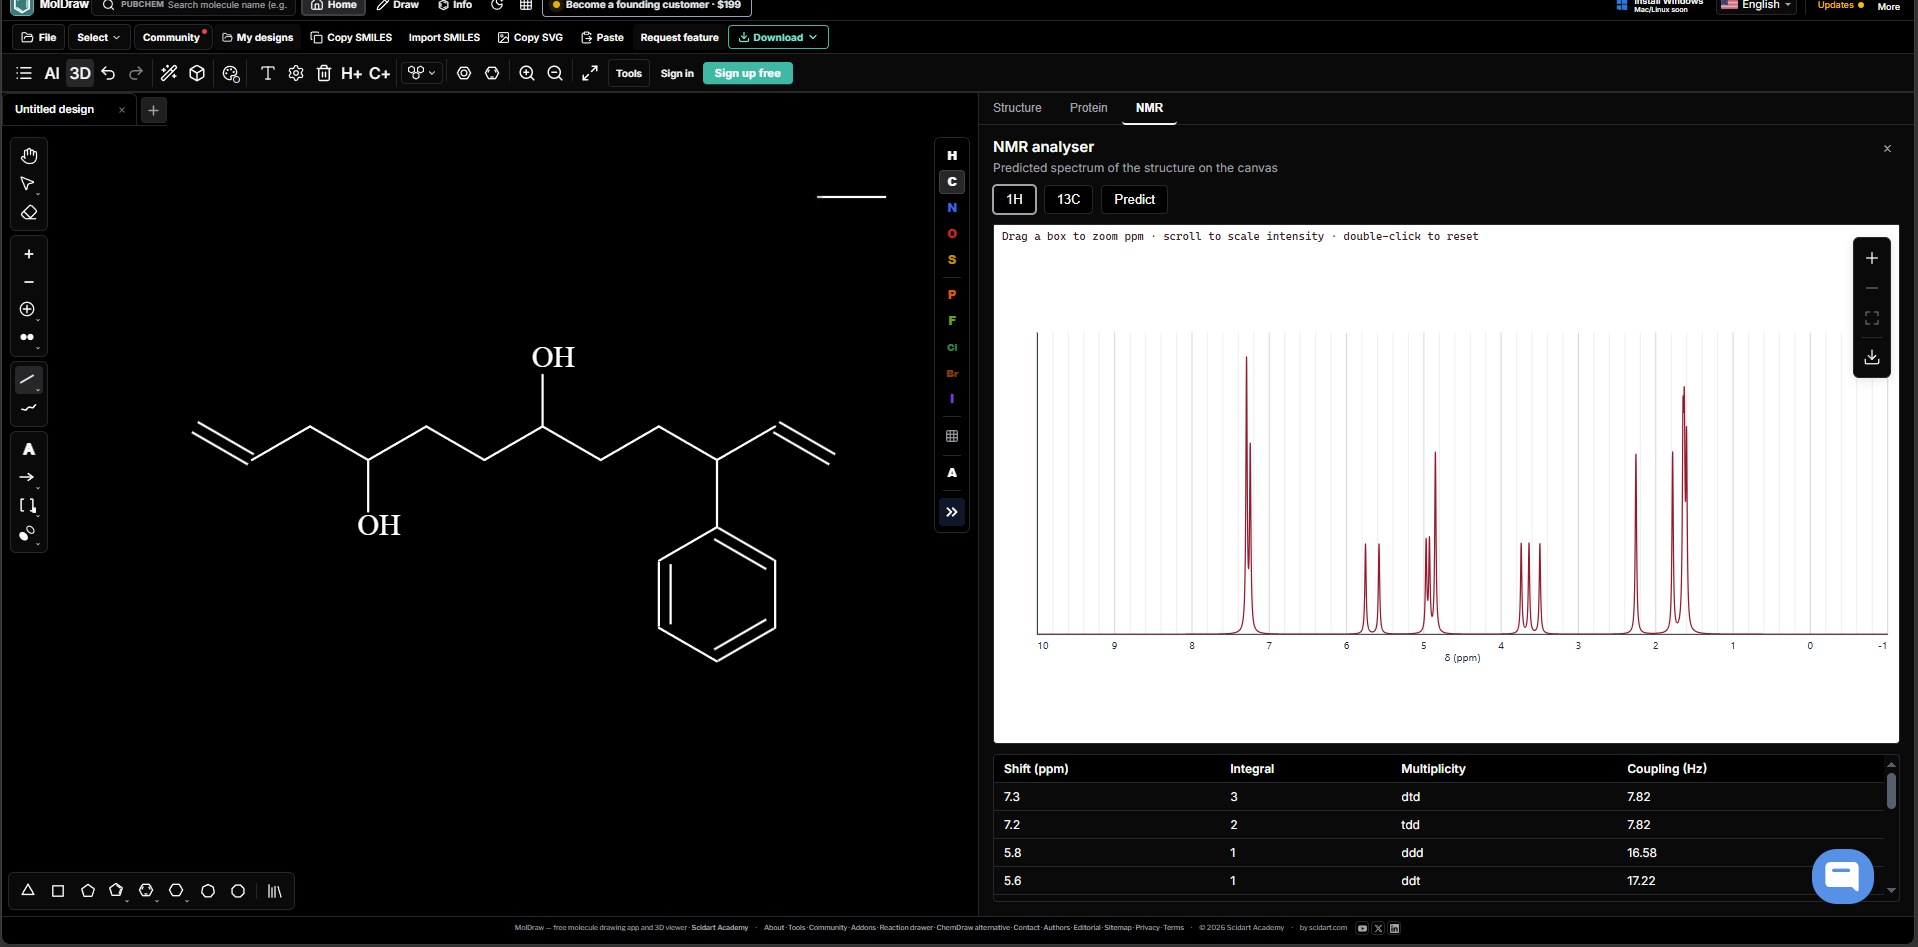
NMR prediction is being developed now. (in beta)
Select a molecule > right click > nmr predict
▲
Select a molecule > right click > nmr predict
Now you can predict NMR spectra of molecules It is an experimental feature. https://www.moldraw.com/tools/free-chem-tools/structure-to-nmr.html
I've been trying to use these two smile codes but it isn't working. Are there any solutions? If there is something I am doing wrong when I click import smiles, let me know. [Na+].OC([O-])=O.Cl>>OC(O)=O.[Na+].[Cl-] [Na+].[Na+].[O-]C([O-])=O.Cl>>[Na+].OC([O-])=O.[Na+].[Cl-]
like to add these type of bond
You can adjust 3D rotation of molecules in Moldraw now!
You can draw nice arrows more easily now in moldraw Try and let me know what to improve.
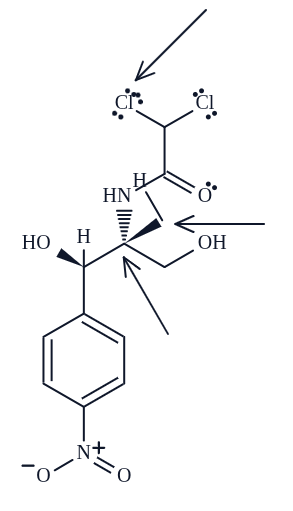
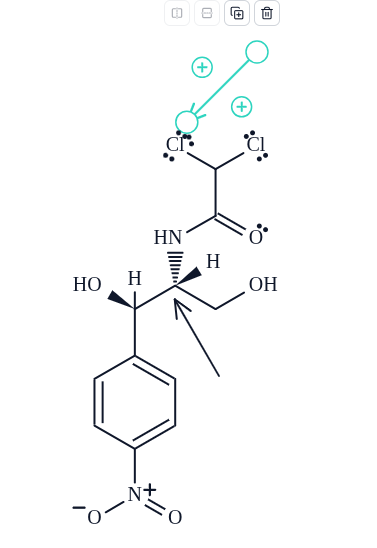
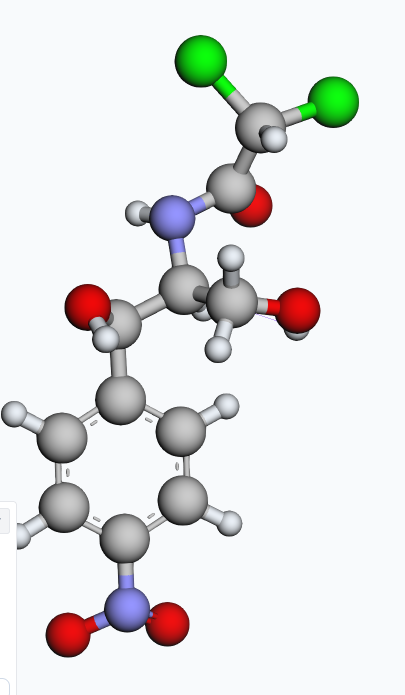
Hi, i really like your update but there is a problem with some things. First i realized now that sometimes editing/deleting posts does not work. In the pictures you can see several problems: - wrong 3d structure at the chiral center when changing the wedge direction - weird LP placement of third LPs (didnt test more than 3) - a button to add one to all of them would be nice so not each atom would have to be clicked - replacing atoms with an h fails and a methyl group is added instead - some lps are drawn wrong when choosing electron flow - in electron flow mode it is not always possible to choose the middle of a bond as endpoint - when a bond is chosen as starting point and an atom as endpoint it often points wrongly which corrects itself when some atoms are deleted (no picture, but could be done on request) Then it is also slightly annoying that enter in the forum results in the post being posted immediately as it is usually used for newlines. It would be nicer if you could only post by clicking the button or have a confirmation before hand. Thanks for your dedication and continued updates. Best Maxim
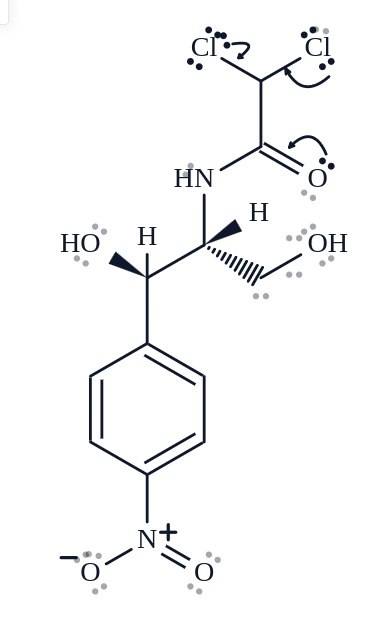
Looks nice but, now I can't dragging atoms or groups to attach them in to an existing structure.
Looks nice but, now I can't dragging atoms or groups to attach them in to an existing structure.
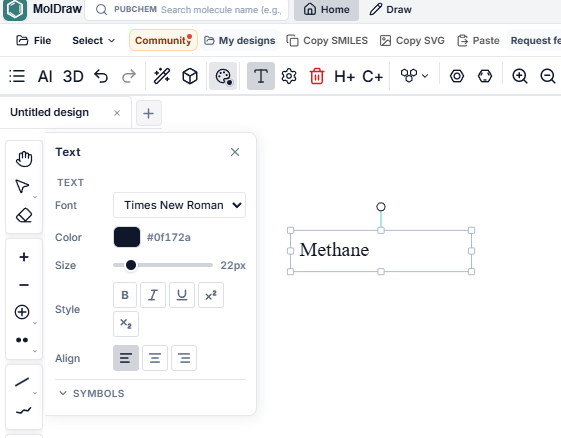
Text tool had some issues. They are fixed. Tell me if anything to be improved further. Thanks.
Select a molecule > right click > nmr predict